
Working standard in pharma: the short answer
A working standard is an in-house lot of a drug substance that a Quality Control laboratory uses in place of the official reference standard for routine testing. Indian GMP calls it a secondary or working standard, and it may be used only after it has been standardised against an official reference standard, initially and at regular intervals thereafter[1].
Working standard qualification is that standardisation: identity by IR, water or loss on drying and assay in triplicate against the primary reference standard, an assigned potency on the as-is basis, and a written statement tracing the lot back to the primary standard it was compared with[2][3]. In India the primary is the IP Reference Substance (IPRS) procured from the Indian Pharmacopoeia Commission; a USP, Ph. Eur. or BP standard may be used where no IPRS exists[2].
Below: the complete SOP of working standard with every limit tagged by its source, what each authority actually requires, a decision tool, a potency calculator and the errors that most often surface in audits.
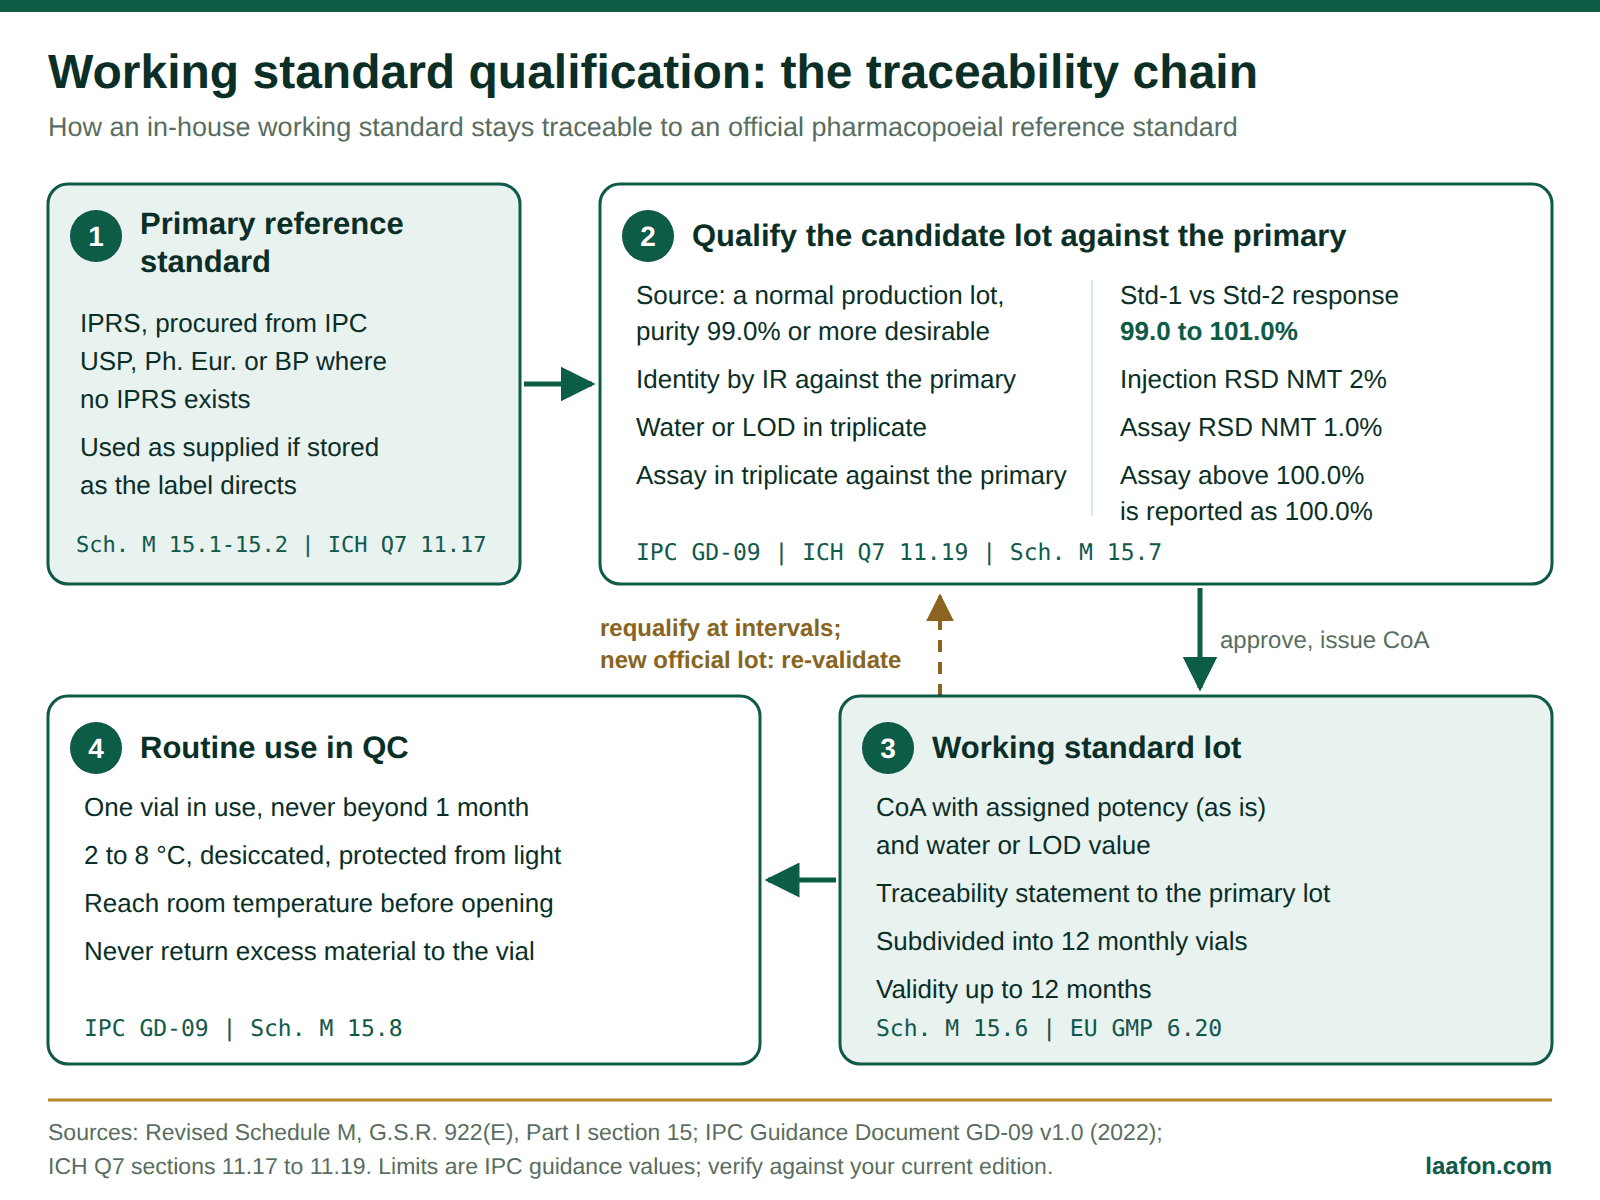
SOP on preparation and handling of working standard
This is the SOP of working standard this page has always carried, rebuilt. The section numbering is kept so an existing adaptation still maps across, but every limit now carries a tag saying where it comes from, and the parts an inspector actually asks about have been added: the traceability statement, intermediate checks, the new-lot rule and what to do when a check fails.
Tag key: statutory Schedule M text IPC guidance IPC GD-09 or ICH/WHO guidance site policy an internal convention you may change check source verify against your edition
1.0 Purpose
To lay down the procedure for selection, qualification against a primary reference standard, subdivision, labelling, storage, use, requalification and disposal of working (secondary) standards used in the Quality Control laboratory.
2.0 Scope
Applies to working standards of active substances, impurities and excipients established in-house for chemical tests in the Quality Control department of [Company name].
It does not cover: official primary reference standards (IPRS, USP, Ph. Eur., BP), which are used as supplied and controlled through the reference standard register; biological and microbiological standards except where stated; volumetric solutions; and standard solutions prepared from a standard for a single analysis.
3.0 Responsibility
- Analyst, QC: sampling, qualification testing, labelling, and entries in the usage record.
- Officer or Executive, QC (standards custodian): register, storage, issue, reconciliation and scheduling of intermediate checks.
- Head, QC: review of qualification data and approval of the working standard certificate of analysis.
- Head, QA: approval of this SOP, review of deviations and periodic review. Accountability rests with Head QA.
4.0 Definitions
- Primary reference standard: a substance whose assigned content is accepted without comparison with another chemical substance[5]. IPRS are primary standards in this sense[2].
- In-house primary standard: established when no official primary exists, after testing that fully establishes identity and purity[3].
- Working (secondary) standard: a substance whose characteristics are assigned by comparison with a primary reference standard[5].
- Standard solution: a solution prepared from a reference or working standard for a particular analysis. Outside the scope of this SOP.
5.0 Procedure
5.1 Selection of batch
5.2 Assigning working standard numbers
CO/WS/XXX/NN/YY-YY, where CO is the company code, WS denotes working standard, XXX the product code, NN the serial number of the working standard in that year, and YY-YY the financial year (for example 26-27). site policy5.3 Qualification against the primary reference standard
5.4 Approval, certificate and packing
5.5 Validity, intermediate checks and requalification
5.6 Storage and handling
5.7 Supply of working standard to customers or other sites
5.8 Deviation handling
5.9 General conditions
6.0 Acceptance criteria
| Parameter | Limit | Basis |
|---|---|---|
| Source material purity (assay use) | 99.0% or more, desirable | IPC guidance anhydrous or volatile-free basis[2] |
| Identification | IR concordant | IPC guidance against primary standard or IP spectrum[2] |
| Water or LOD, related substances | monograph limits | check source current monograph for the substance |
| Std-1 vs Std-2 response correlation | 99.0–101.0% | IPC guidance[2] |
| Replicate and bracketing standard injections, overall %RSD | NMT 2% | IPC guidance[2] |
| Assay, %RSD of triplicate | NMT 1.0% (micro: 5.0%) | IPC guidance[2] |
| Assay reported above 100.0% w/w | report as 100.0% | IPC guidance not for biological standards[2] |
| Agreement between analysts | NMT 0.5% absolute | site policy not a compendial or IPC requirement |
| Water or LOD agreement | absolute difference | site policy set from method precision data, not %RSD |
| Validity of the lot | up to 12 months | IPC guidance extendable on data[2] |
| Use period of an opened vial | NMT 1 month | IPC guidance[2] |
| Standardisation against official standard | initially and at intervals | statutory Schedule M 15.7[1] |
Swipe the table sideways on a phone. IPC values come from a guidance document, not the monograph; confirm them against the current IP edition and your approved specification.
7.0 Frequency
| Activity | When | Basis |
|---|---|---|
| Qualification of a new lot | before first use | ICH Q7 11.19[3] |
| Intermediate check (water or LOD, purity, assay) | on a written schedule, for example at mid-validity | site policy requirement to check is IPC guidance[2] |
| Re-validation | new official primary lot, or major change in the monograph test | IPC guidance[2] |
| Change of in-use vial | monthly, or earlier if consumed | IPC guidance[2] |
| Desiccant regeneration | at indicator colour change | site policy |
8.0 Precautions
- Keep vials tightly closed and upright; never open a cold vial.
- Never dispense directly from the vial into a volumetric flask, and never return material.
- Record every weighing contemporaneously; qualification raw data are subject to the same ALCOA+ data integrity expectations as release data.
- Do not use a working standard in a dispute or referee analysis; use the official reference standard.
9.0 Annexures
- Annexure-I: Working standard qualification record (format below)
- Annexure-II: Working standard consumption and reconciliation record
- Annexure-III: Desiccant regeneration and replacement record
- Annexure-IV: Specimen label for working standard vial
Annexure-I: Working standard qualification record
| Test | Det. 1 | Det. 2 | Det. 3 | Mean | %RSD or diff. | Limit | Done by | Checked by |
|---|---|---|---|---|---|---|---|---|
| Primary standard name, lot, valid-use status | current lot | |||||||
| Identification (IR) | concordant | |||||||
| Water or LOD (%) | monograph | |||||||
| Std-1 vs Std-2 correlation (%) | 99.0-101.0 | |||||||
| Assay, dried or anhydrous basis (%) | RSD NMT 1.0 | |||||||
| Assigned potency, as is (%) | max 100.0 |
Copies as tab-separated text that pastes straight into Excel or Word.
10.0 Revision history
| Version | Effective date | Change | Change control no. |
|---|---|---|---|
| 00 | DD-MMM-YYYY | New SOP | ___ |
| 01 | DD-MMM-YYYY | Aligned with revised Schedule M Part I section 15 and IPC GD-09: traceability statement, intermediate checks, new-lot rule, deviation handling | ___ |
This SOP is a template for adaptation. It requires local qualification, validation and Quality Assurance approval before use, and every acceptance criterion must be verified against the current pharmacopoeial edition and your approved specifications. Pharmacopoeial texts, IPC guidance and Indian statutory instruments change between editions.
Working standard qualification as per IP, USP, ICH, Ph. Eur. and Schedule M
Searches for working standard qualification “as per USP” or “as per ICH” assume each body publishes its own procedure. Most do not. The tabs below show what each one actually says, and what it leaves to you.
Indian Pharmacopoeia Commission: GD-09 (2022)
The most detailed public text on the subject, and the one most Indian SOPs have never cited. IPC’s guidance document on IP Reference Substances, version 1.0 of 1 June 2022, has a full chapter on working standards[2]. It:
- defines a secondary standard, “commonly known as working standard”, as usable once qualified against a primary standard, with IPRS as the primary and USP, Ph. Eur. or BP standards acceptable where no IPRS is available;
- requires IR identity, and water or LOD plus assay in triplicate against IPRS;
- sets the injection sequence, the 99.0–101.0% standard correlation, NMT 2% injection RSD, NMT 1.0% assay RSD and the 100.0% reporting cap;
- allows validity up to 12 months with extension on data, one month per opened vial, intermediate checks, and re-validation against every new official IPRS lot;
- specifies label contents, Type I amber glass and suitable rubber closures, and forbids returning excess material to the vial.
What it does not say: it does not require three analysts, and it sets no limit on agreement between analysts.
Revised Schedule M, Part I, section 15
Notified as G.S.R. 922(E) on 28 December 2023, section 15 “Reference standards” is the statutory text in India[1]:
- 15.1 official reference standards shall be used whenever they exist; 15.2 IP reference standards shall be procured from the Indian Pharmacopoeia Commission;
- 15.3 official standards are used only for the purpose described in the monograph; 15.4 standards prepared by the manufacturer are tested, released and stored like official ones;
- 15.5 secondary or working standards may be established by appropriate tests and checks at regular intervals;
- 15.6 labels carry at least name, batch or lot and control number, date of preparation, shelf life, potency and storage conditions;
- 15.7 all in-house working or secondary standards shall be standardised against an official reference standard, when available, initially and at regular intervals thereafter;
- 15.8 all reference standards are stored and used so their quality is not affected.
What it does not say: no numeric limits. Those come from IPC GD-09 and your specification.
ICH Q7, sections 11.17 to 11.19
Written for API manufacture (Step 4, November 2000) and often used as the reference text by finished-product laboratories as well[3]:
- 11.17 the source of each primary standard is documented; standards from an officially recognised source are normally used without testing if stored as the supplier recommends;
- 11.18 where no official primary exists, an in-house primary standard is established with testing that fully establishes identity and purity;
- 11.19 each batch of secondary standard is prepared, identified, tested, approved and stored appropriately, its suitability determined before first use by comparison with a primary, and it is periodically requalified to a written protocol.
What it does not say: ICH Q7 names no tests, replicate counts, RSD limits or validity period. “As per ICH” means the three obligations above, with the numbers set in your protocol.
EU GMP Chapter 6 and Ph. Eur. 5.12
EU GMP Part I, clause 6.20 (in operation from 1 October 2014) says compendial reference standards should preferably be used as primary standards, and that secondary standards are permitted once their traceability to primary standards has been demonstrated and documented. Clause 6.21 requires reference standards to be marked with the preparation and opening dates and the signature of the person who prepared them[4].
Ph. Eur. general chapter 5.12 defines the secondary standard. Its text is not freely available, but EDQM and USP scientists quoted it in a joint 2023 presentation: a secondary standard “should exhibit the same property or properties as the primary standard” for the tests it is established for. The same presentation recommended showing equivalence of results by statistical tests and noted that results obtained with the pharmacopoeial standard alone are conclusive[9].
What was not verified: the full text of Ph. Eur. 5.12 was not retrieved for this page, so no limit on this page is attributed to it.
WHO: TRS 943 Annex 3 and TRS 1052 Annex 4
WHO’s general guidelines on chemical reference substances (2007) define primary and secondary substances, and note that a manufacturer’s working standards can in principle be prepared by the same procedures. Where an assay standard is expressed “as is” and the assay is non-specific, its water and residual solvent content must be determined; storage at about 5 °C has proved satisfactory for most substances; single-use units are preferred, and multi-use containers need more frequent re-testing[5].
WHO’s 2024 good practices for QC laboratories (clauses 5.74 to 5.81) require pharmacopoeial substances where available, allow manufacturers to use secondary (working) standards traceable to primary ones, and require an identification number per batch quoted on worksheets, a register, confirmation of validity before use, and retesting at regular intervals with a retrospective check if a standard fails[6].
Status note: WHO circulated a draft revision of the reference substances guideline (QAS/25.972) for comment in 2025[10]. At the time of writing the 2007 text was the published version found; check for an adopted revision.
USP: there is no “as per USP” procedure
USP publishes no working standard qualification procedure. Its reference standards FAQ states plainly that USP does not provide guidance on the qualification or use of non-USP secondary reference standards, including in-house standards, and that a USP Reference Standard explicitly mentioned in an official procedure is part of the official method[7].
In practice “qualification as per USP” therefore means: qualify against the current USP lot; confirm before each use that the lot is current or a previous lot within its valid-use date, which USP makes the user’s responsibility; and treat results obtained with the USP standard as the reference if a result is ever disputed[7].
US FDA: 21 CFR 211.194(c)
The US regulation is short: complete records shall be maintained of any testing and standardisation of laboratory reference standards, reagents and standard solutions[8]. It sets no procedure and no numbers. An FDA inspector will read your qualification record against your own SOP, which is why the site-policy rows in section 6.0 need a written justification.
The pattern across all seven: the obligation to qualify against a primary standard, document traceability and requalify is universal. The numbers are not. In India, IPC GD-09 is the only public source of numeric limits; everything else in a typical SOP, including three analysts and the 0.5% rule, is a site convention that should be labelled as one.
Which standard should you use? Decision tool
Two questions decide what the procedure has to do. Choose one answer in each group.
1. Is an official pharmacopoeial reference standard available for this substance?
2. What are you doing?
Choose one option from each group.
The verdict and the clause it rests on will appear here.
Assigned potency calculator
Enter the three assay results on the dried or anhydrous basis and the three water or LOD results. The calculator returns the mean, the assay %RSD against the IPC limit, the water spread, and the assigned potency on the as-is basis that goes on the certificate and the vial label.
Example values loaded. Press Calculate.
- Mean assay, dried basis–
- Assay %RSD (IPC limit NMT 1.0%)–
- Mean water or LOD–
- Water spread, highest minus lowest–
- Assigned potency, as is–
As is = dried-basis assay × (100 − water or LOD) ÷ 100. A dried-basis mean above 100.0% is capped at 100.0% before conversion, following IPC GD-09. IPC does not state which basis the cap applies to; this calculator applies it to the dried basis, so match your own SOP.
The sample RSD uses n minus 1. Nothing you enter leaves your browser. The calculator checks arithmetic only; it does not replace the reviewed qualification record.
Six errors in older working standard SOPs
Each of these appeared in the earlier version of this page, and each appears in many SOPs still in use. They are the first things a knowledgeable auditor checks.
- Naming the wrong source for official standards. Older SOPs send the analyst to Central Drugs Laboratory, Kolkata. Revised Schedule M 15.2 now says IP reference standards shall be procured from the Indian Pharmacopoeia Commission[1].
- An RSD limit on water content. A 1.0% RSD limit for water or LOD is unachievable at low levels: three good results of 0.19, 0.20 and 0.21% give 5% RSD. Use an absolute difference based on method precision.
- Presenting site conventions as regulation. Three analysts and a 0.5% inter-analyst limit are reasonable choices, but neither IPC GD-09 nor ICH Q7 requires them[2][3]. Label them as site policy so an inspector does not read them as a misquoted standard.
- No trigger for a new primary lot. When a new IPRS lot becomes official, the working standard is re-validated against it[2]. An SOP that only requalifies annually misses this.
- No traceability statement. A certificate that gives a potency but not the primary standard and lot it was assigned against does not demonstrate traceability, which EU GMP 6.20 requires to be documented[4].
- Conflicting vial numbering and no failure path. Numbering monthly vials 1/13 to 13/13 while calling the reserve 1/1 invites mix-ups, and an SOP with no deviation section says nothing about the retrospective review a failed check demands[2][6].
Working standard vs reference standard vs working standard solution
Three terms that are often used interchangeably, and should not be.
| Aspect | Primary reference standard | Working standard | Standard solution |
|---|---|---|---|
| What it is | Official substance (IPRS, USP, Ph. Eur., BP) or a fully characterised in-house primary | In-house lot qualified against the primary | A solution prepared from either, for one analysis |
| Qualified how | By the issuing body; used as supplied[3] | IR, water or LOD and assay in triplicate against the primary[2] | Prepared as the method directs; not qualified separately |
| Validity | While the lot is current, or within the valid-use date of a previous lot[7] | Up to 12 months, one month per opened vial[2] | As stated in the method or solution stability data |
| Typical use | Qualifying working standards; dispute and referee testing | Routine release and stability testing | Injection or measurement in that run |
| Indian hook | Schedule M 15.1 to 15.3[1] | Schedule M 15.5 and 15.7[1] | Laboratory reagent and solution controls |
Frequently asked questions
A working standard is an in-house lot of a drug substance qualified against an official primary reference standard, such as an IPRS, and then used for routine QC testing in its place. Revised Schedule M calls it a secondary or working standard and requires it to be standardised against an official reference standard initially and at regular intervals.
USP does not publish one. Its reference standards FAQ says USP does not provide guidance on qualifying non-USP secondary standards, including in-house standards. Laboratories qualify against the current USP lot using the GMP obligations in ICH Q7 11.19 and their own written protocol, and treat results with the USP standard as the reference in any dispute.
IPC guidance document GD-09 allows validity of up to twelve months, extendable where stability, storage and extent of use justify it, and shorter for hygroscopic or unstable materials. An individual opened vial should not be used beyond one month. Many sites also cap validity at the retest date of the source batch as internal policy.
IPC gives twelve as its example, one vial for each month of validity, each holding about one gram or a suitable quantity. Many laboratories add one sealed reserve vial for investigations and customer supply; that thirteenth vial is site policy, not an IPC requirement.
Yes. IPC GD-09 states that whenever a new lot of the primary standard becomes official, the working standard should be prepared or re-validated only against the new lot. Build this trigger into the SOP alongside the scheduled intermediate checks.
IPC GD-09 says an assay above 100.0% w/w is considered and reported as 100.0% w/w, except for biological standards. A result well above 100% is also a prompt to check the primary standard’s potency value, the dilution scheme and the water correction before accepting the lot.
No. A working standard is a qualified solid lot with its own certificate and validity. A standard solution is prepared from a reference or working standard for one analysis, and its stability is set by the analytical method, not by the working standard SOP.
No. Schedule M requires official reference standards to be used whenever they exist and only for the purpose in the monograph, and results obtained with the pharmacopoeial standard are the conclusive ones. Keep the official standard for qualifying working standards and for any dispute or referee testing, and consider it when confirming an out-of-specification result.
Preparing the QC laboratory for a Schedule M inspection?
Reference and working standard control is one of the systems we review during a Revised Schedule M gap assessment, alongside SOPs, qualification records and laboratory data integrity. The work is done by practitioners with more than two decades in pharmaceutical QA, QC and regulatory affairs.
Related on Laafon
References
- Ministry of Health and Family Welfare, Government of India. Drugs Rules, 1945: Schedule M, Good Manufacturing Practices and Requirements of Premises, Plant and Equipment for Pharmaceutical Products. G.S.R. 922(E), 28 December 2023. Part I, section 15, Reference standards. New Delhi: CDSCO; 2023. Available from: https://cdsco.gov.in/opencms/opencms/system/modules/CDSCO.WEB/elements/download_file_division.jsp?num_id=MTA4MTU%3D. Accessed September 2026.
- Indian Pharmacopoeia Commission. Guidance document: Indian Pharmacopoeia Reference Substances. IPC/GD/09, version 1.0. Ghaziabad: IPC; 1 June 2022. Sections on working standards, validity, intermediate checks, labels and packaging. Available from: https://www.ipc.gov.in/images/GD-09-IP_Reference_Substances.pdf. Accessed September 2026.
- International Council for Harmonisation. ICH Q7: Good Manufacturing Practice Guide for Active Pharmaceutical Ingredients. Step 4, 10 November 2000. Sections 11.17 to 11.19. Geneva: ICH; 2000. Available from: https://database.ich.org/sites/default/files/Q7%20Guideline.pdf. Accessed September 2026.
- European Commission. EudraLex Volume 4, EU Guidelines for Good Manufacturing Practice, Part I, Chapter 6: Quality Control. Clauses 6.20 and 6.21. Brussels: European Commission; in operation from 1 October 2014. Available from: https://health.ec.europa.eu/system/files/2016-11/2014-11_vol4_chapter_6_0.pdf. Accessed September 2026.
- World Health Organization. General guidelines for the establishment, maintenance and distribution of chemical reference substances. WHO Technical Report Series No. 943, Annex 3. Geneva: WHO; 2007. Available from: https://www.who.int/docs/default-source/medicines/norms-and-standards/guidelines/quality-control/trs943-annex3-establishmentmaintenance-distribution-chemica-reference-substances.pdf. Accessed September 2026.
- World Health Organization. WHO good practices for pharmaceutical quality control laboratories. WHO Technical Report Series No. 1052, Annex 4. Geneva: WHO; 2024. Clauses 5.74 to 5.81. Available from: https://www.who.int/publications/m/item/who-good-practices-for-pharmaceutical-quality-control-laboratories. Accessed September 2026.
- United States Pharmacopeia. Reference Standards: frequently asked questions (Uses and Development; Availability and Validity). Rockville (MD): USP. Available from: https://www.usp.org/frequently-asked-questions/reference-standards. Accessed September 2026.
- US Food and Drug Administration. Code of Federal Regulations, Title 21, section 211.194(c), Laboratory records. Washington (DC): eCFR. Available from: https://www.ecfr.gov/current/title-21/chapter-I/subchapter-C/part-211/subpart-J/section-211.194. Accessed September 2026.
- Almeling S, Zeine C. Secondary standards: considerations in traceability to pharmacopoeial standards [presentation]. Joint EDQM and USP session, 10 October 2023. Strasbourg: EDQM; 2023. Available from: https://www.edqm.eu/documents/52006/1817824/Presentation+-415+Secondary+Standards…. Accessed September 2026.
- World Health Organization. Working document QAS/25.972: General guidelines for the establishment, maintenance and distribution of chemical reference substances, draft revision for comment. Geneva: WHO; February 2025. Available from: https://cdn.who.int/media/docs/default-source/medicines/norms-and-standards/current-projects/2025-02-25-referencesubstances-qas25-972.pdf. Accessed September 2026.
Technical and educational content only, not legal or regulatory advice. The SOP above is a template: it requires local qualification, validation and Quality Assurance approval before use. Acceptance criteria must be verified against the current pharmacopoeial edition, the current IPC guidance and your approved specifications; pharmacopoeial texts and Indian statutory instruments change between editions. Reviewed by Darshan Singh, 23+ years in pharmaceutical QA, QC and regulatory affairs. Last reviewed 11 September 2026.