
A control sample of finished product must be kept for at least one year after the expiry date, in its final packaging, in a quantity sufficient for at least two full re-examinations. All three of India, the EU and the US land on that same one-year figure — Schedule M clause 19.8.2, EU GMP Annex 19 clause 3.1, and 21 CFR 211.170(b)(1) [1][2][4].
Where published control sample guidelines go wrong is everything after that sentence. Starting materials, active starting materials and API reserve samples run on three different clocks anchored to three different events, only one regulator excludes sterility and pyrogen testing from the quantity calculation, and the term “control sample” does not appear anywhere in India’s notified Schedule M.
Control sample guidelines: what the three regulators actually require
Three instruments govern retained samples for most Indian manufacturers: the revised Schedule M to the Drugs Rules, 1945, notified by G.S.R. 922(E) on 28 December 2023 [1]; EU GMP Annex 19, for anyone exporting to or manufacturing for the European market [2]; and 21 CFR 211.170 for the United States [4]. They agree on the headline number for finished product and diverge on nearly everything else.
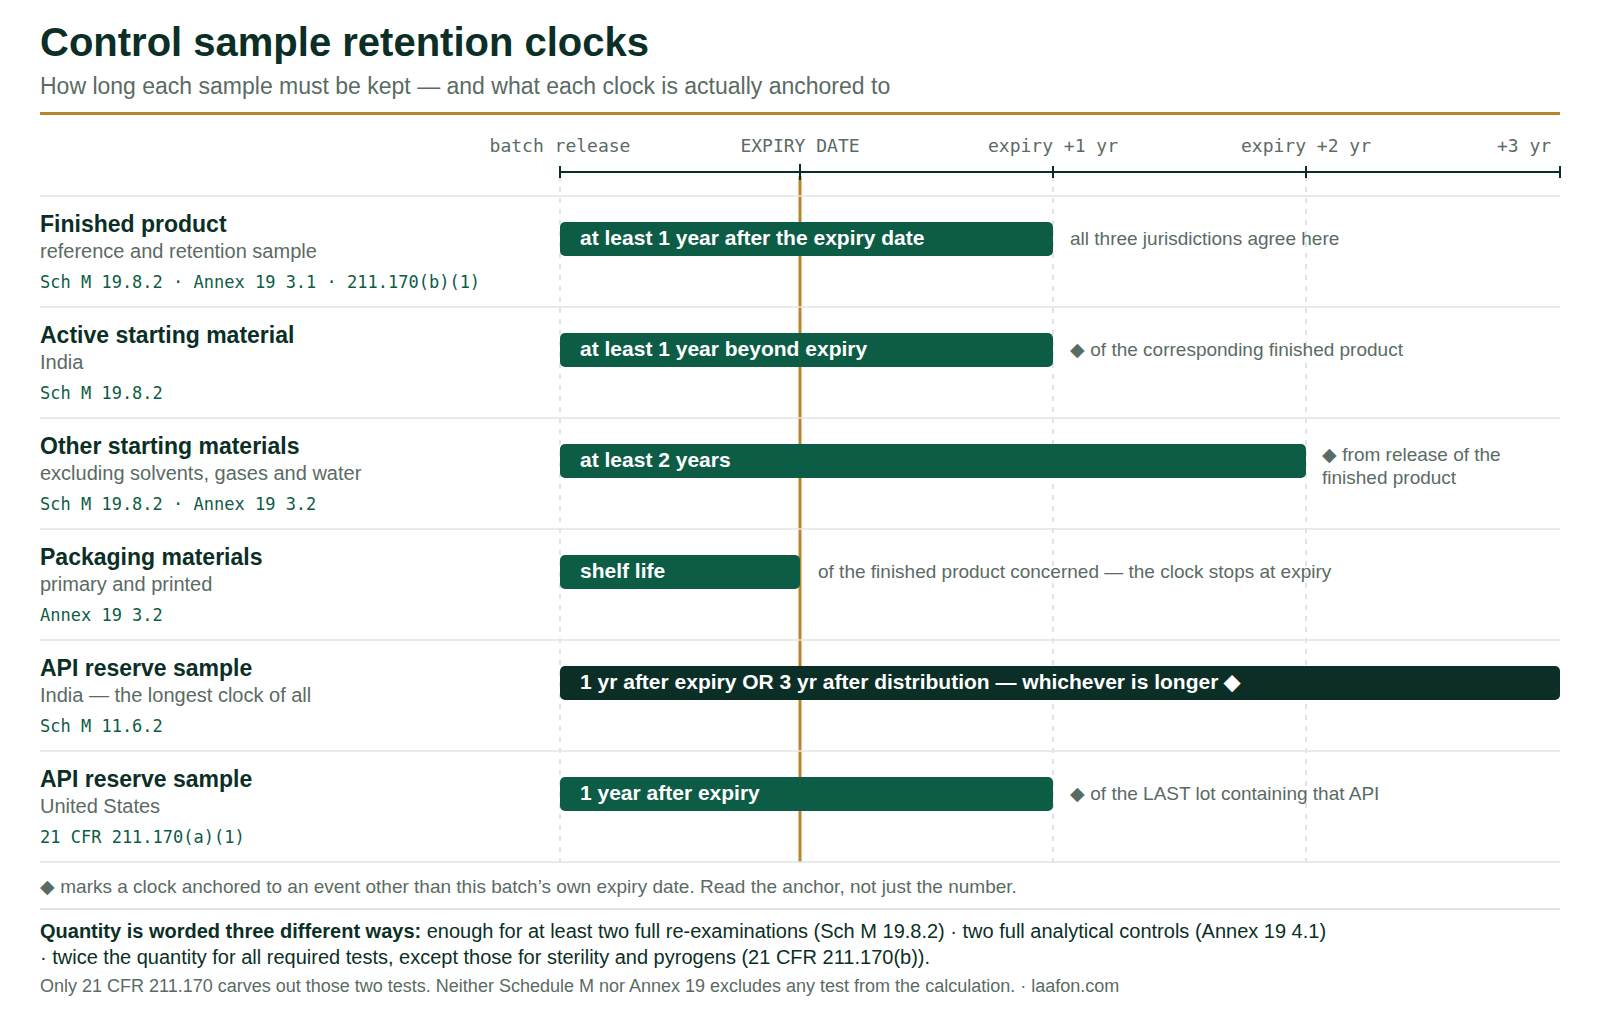
| Sample | India — Schedule M | EU — Annex 19 | US — 21 CFR 211.170 |
|---|---|---|---|
| Finished product | At least 1 year after the expiry date 19.8.2 | At least 1 year after the expiry date 3.1 | 1 year after the expiration date of the drug product 211.170(b)(1) |
| Active starting material | At least 1 year beyond the expiry date of the corresponding finished product 19.8.2 | Treated as a starting material: at least 2 years after release of the finished product 3.2 | 1 year after the expiration date of the last lot of drug product containing it 211.170(a)(1) |
| Other starting materials excl. solvents, gases, water |
Minimum 2 years, if their stability allows 19.8.2 | At least 2 years after release of the finished product; may be shortened if the specification stability period is shorter 3.2 | Not addressed — 211.170 covers active ingredients and drug products only |
| Packaging materials | Not separately specified in Part I clause 19.8.2 | For the duration of the shelf life of the finished product concerned 3.2 | Not addressed by 211.170 |
| API reserve sample at the API maker |
1 year after expiry or 3 years after distribution of the batch, whichever is longer Part XII 11.6.2 | Not in Annex 19 — API manufacture sits under EU GMP Part II | 1 year after the expiration date of the last lot containing the active ingredient 211.170(a)(1) |
| Quantity | Sufficient for at least two full re-examinations 19.8.2 | Sufficient for two occasions of the full analytical controls in the MA file. No test is excluded 4.1 | Twice the quantity for all required tests, except those for sterility and pyrogens 211.170(b) |
| Pack | Usually the final packaging; smaller samples in appropriate containers where packages are exceptionally large 19.8.2, 19.3(g) | Finished primary packaging, or packaging of the same material as the marketed primary container 3.1 | The same immediate container-closure system as marketed, or one with essentially the same characteristics 211.170(b) |
Swipe the table sideways on a phone. Clause numbers refer to the instrument named in that column’s heading.
The exclusion most Indian SOPs misattribute. The phrase “twice the quantity required to conduct all the tests except sterility and pyrogens / bacterial endotoxin test” appears in thousands of Indian control sample SOPs, usually presented as a GMP or Schedule M requirement. It is neither. That wording is 21 CFR 211.170, a United States regulation [4]. Searching the full notified text of Schedule M returns no occurrence of “twice the quantity”, and EU GMP Annex 19 clause 4.1 excludes no test at all from its two-full-controls calculation. If your SOP carries that sentence and you do not export to the US, you are citing the wrong instrument.
The practical consequence is not academic. Under Annex 19 there is no sterility or pyrogen carve-out, so a sterile product’s reference sample must be sized for two complete sets of controls including those tests. A site that sized its retention quantity from a US-derived SOP while supplying the EU market can find itself short of sample at exactly the moment a complaint investigation needs it.
Why “control sample” is not a regulatory term
“Control sample” is Indian industry vocabulary. It is not the vocabulary of the regulation. A character search of the complete notified Schedule M — all thirteen Parts — returns zero occurrences of “control sample” or “counter sample”. The notified text says retention samples (Part I clause 19.8.2), reference samples (Part I clause 9.5.6, in the loan licence and contract context) and reserve samples (Part XII clause 11.6, for APIs) [1].
That matters because the two principal terms are not synonyms. EU GMP Annex 19 clause 2.1 defines them as answering two different purposes [2]:
| Term | What it is | What it is kept for | Source |
|---|---|---|---|
| Reference sample | A sample of a batch of starting material, packaging material or finished product | To be analysed, should the need arise during the shelf life of the batch | Annex 19 2.1 |
| Retention sample | A sample of a fully packaged unit from a batch of finished product | Identification — presentation, packaging, labelling, leaflet, batch number, expiry date | Annex 19 2.1 |
| Reserve sample | The US term, covering both active ingredients and drug products | Potential future evaluation of the quality of the batch | 21 CFR 211.170 |
Annex 19 adds the concession that for finished products the two “will be presented identically, i.e. as fully packaged units”, and in those circumstances “may be regarded as interchangeable”. That concession is why the Indian habit of collapsing both into one “control sample” usually survives an inspection — but it is a concession, not a definition, and it does not hold where the reference sample is drawn at an intermediate stage or where the packaged unit is too large to retain whole.
Schedule M does draw the distinction — in exactly one place. Part IV, which covers the manufacture of biological products, states at clause 13.2 that samples for post-release use “typically fall into one of two categories reference samples or retention samples for the purposes of analytical testing and identification respectively”, then sets separate periods at 13.2.1 and 13.2.2 [1]. Part I, which governs everything else, does not carry that definition. So an Indian biologicals manufacturer is held to the Annex 19 two-category scheme by name, and a tablet manufacturer is not.
Retention clock and disposal date calculator
Pick the jurisdiction and the sample type. The clause and the rule update together; enter the dates and the earliest permissible disposal date is calculated from that rule.
Choose a jurisdiction and a sample type
Clause reference appears here
Enter the dates to calculate the earliest disposal date.
The calculator applies the minimum period stated in the instrument. Your registered dossier, a national law under Annex 19 clause 3.2, or your own quality agreement may require longer — the longer period governs.
The root-n-plus-one formula and what it does not cover
Many Indian control sample SOPs — including the earlier version of this page — instruct that samples be collected “on the basis of formula root-n plus one”. Two things are worth knowing before that line goes into a document an inspector will read.
First, it is not a retention sample rule. The square root of n plus one is a container-selection heuristic for incoming material sampling. It says nothing about how many units of finished product to retain, and no instrument applies it to retention quantity. Schedule M sets retention quantity by a different standard entirely: sufficient for at least two full re-examinations [1].
Second, the formula has no compendial or statistical pedigree. Saranadasa, writing in Pharmaceutical Technology, traces it to a 1920s need to give agricultural regulatory inspectors a convenient, memorisable rule; it was formalised by an Association of Official Agricultural Chemists committee in 1927 for certain classes of foods [6]. Objections are as old as the rule itself, and Borland’s 1950 paper is titled, without hedging, “The Fallacy of Square Root Sampling Rule”. Saranadasa’s own conclusion is that the rule “should not be used to select a sampling plan to infer a population defective rate”.
The numbers make the point concretely. For a lot of 100 containers the formula gives a sample of 11. Accepting on zero defectives, that sample carries a Type I error of 25.8% against a 10% defect level, 48.7% against 5%, and 79.1% against 1%. Reaching 95% confidence at those defect levels would require 23, 39 and 78 containers respectively [6].
What Schedule M requires instead. Clause 19.7.2 states that an identity test shall be conducted on a sample from each container of starting material, and that sampling only a proportion of containers is permissible where a validated procedure has been established to ensure that no single container has been incorrectly labelled. Clause 19.7.2.1 lists what that validation must take account of. A character search of the notified Schedule M finds no occurrence of a square-root sampling formula in any form [1]. The rule of thumb is not prohibited, but it is not the requirement, and it cannot substitute for the validation that clause 19.7.2 actually asks for.
Control sample room: storage, register and the records that fail audits
Schedule M clause 19.8.2 requires finished product retention samples to be kept “in their final packaging and stored under the recommended conditions”. Annex 19 clause 5.2 requires storage in accordance with the marketing authorisation, including refrigerated storage where relevant [2]. Neither instrument states a temperature or a humidity figure for the retention area itself — the condition is inherited from the product’s own labelled storage condition, not set independently.
That is a meaningful negative. A control sample room specification of 25 degrees C and 60% RH is a site decision derived from the products stored in it, and it should be written into your SOP as such rather than presented as a compendial requirement. Where products with different labelled conditions are retained, the room must satisfy each of them, which in practice means segregated areas rather than one averaged set point. Mapping and monitoring of that area follows the same logic as a stability chamber qualification.
On records, 21 CFR 211.170(b) carries a requirement the Indian and EU texts do not: reserve samples from representative lots or batches, selected by acceptable statistical procedures, must be examined visually at least once a year for evidence of deterioration, unless visual examination would affect sample integrity. Any evidence of deterioration must be investigated under 21 CFR 211.192, and the results recorded and maintained with the product’s other stability data [4]. Radioactive drug products are carved out of that duty.
A note on the sample register. Neither Schedule M nor Annex 19 prescribes the format of a control sample register, and Schedule M contains no clause requiring one by that name. Annex 19 clause 2.4 does require that “records of traceability of samples should be maintained and be available for review by competent authorities”. The register in the template below is therefore a site document built to satisfy that traceability obligation — and, being a GxP record, it should meet the same ALCOA+ attributes as any other quality record.
Revised Annex 19 applies from 24 September 2026
The European Commission published a revised Annex 19 on 23 June 2026 as C(2026) 4135 final, applicable three months from publication — 24 September 2026 [3][7]. Before you rewrite an SOP around it, note precisely how narrow it is.
What did not change. The one-year finished product period (3.1), the two-year starting material period (3.2), the packaging material shelf-life rule (3.2) and the two-full-analytical-controls quantity rule (4.1) are carried over unchanged. The reference and retention sample definitions in 2.1 are unchanged. If your product is not parallel traded, the revision does not alter your retention periods at all.
What changed. The stated reason for the revision is reference and retention samples for parallel imported, parallel distributed and parallel traded products, and section 9 is rewritten accordingly [3]:
- 9.2 now states plainly that reference samples of the re-packaged product are not required.
- 9.3 requires a retention sample of the re-packaged finished product for each re-packaging operation, retained at least one year after expiry.
- 9.4 requires that sample to include the primary and the secondary package; where the secondary package is not opened, only the packaging material used need be retained.
- 9.5 is the substantive new permission: where it is duly justified that a physical retention sample cannot reasonably be retained, and agreed in advance with the competent authority, a photographic or digital sample may be retained instead.
- 9.5.1 to 9.5.4 set what that digital sample must carry: a complete record permitting full visual examination, high-quality photographs traceable to the batch packaging record, all data on both primary and secondary packaging, evidence that safety features have been applied, identification of braille information, and compliance with Annex 11 for record integrity over the retention period.
A digital retention sample is a genuine first for EU GMP, and the conditions attached to it are strict: prior agreement with the competent authority, and Annex 11 data integrity controls for the whole retention period. It is a relief for parallel distributors handling cold-chain or high-value packs, not a general licence to photograph retention samples instead of keeping them.
SOP for control sample handling (adaptable template)
1. Purpose
To define the procedure for drawing, labelling, storing, tracking, re-examining and destroying reference and retention samples of starting materials, packaging materials and finished products.
2. Scope
Applies to the Quality Control department at the manufacturing site for all commercial batches of finished product, all starting materials other than solvents, gases and water used in the process, and all primary and printed packaging materials. This procedure does not cover stability study samples, in-process samples, investigational medicinal products, or reserve samples held by an API manufacturer under Schedule M Part XII clause 11.6 — each of those is governed by its own procedure.
3. Responsibility
- QC Chemist / Sampler: drawing, labelling and sealing samples; entering them in the register.
- Officer, Quality Control: verifying quantity against the two-re-examination rule; storage condition monitoring; annual visual examination where applicable.
- Head, Quality Assurance: approval of this SOP, authorisation of destruction, and review of any deviation.
4. Materials and equipment
- Sample containers of the marketed primary packaging, or of the same material as the marketed primary container.
- Low-density polyethylene bags with an inner label, heat sealer, tamper-evident seals.
- Calibrated temperature and humidity monitoring for the retention area, with alarm and data review.
- Retention sample register (Annexure-I) and destruction record (Annexure-II).
5. Procedure
6. Acceptance criteria
| Parameter | Requirement | Basis |
|---|---|---|
| Retention period, finished product | At least 1 year after expiry | statutory Schedule M 19.8.2 [1] |
| Retention period, active starting material | At least 1 year beyond FP expiry | statutory Schedule M 19.8.2 [1] |
| Retention period, other starting materials | Minimum 2 years | statutory Schedule M 19.8.2 [1] |
| Quantity retained | Two full re-examinations | statutory Schedule M 19.8.2 [1] |
| Quantity retained, EU supply | Two full analytical controls, no exclusions | guideline Annex 19 4.1 [2] |
| Quantity retained, US supply | Twice all required tests, less sterility and pyrogens | statutory 21 CFR 211.170(b) [4] |
| Storage condition of the retention area | Per product label / MA | site policy derived from the products held; no compendial figure exists |
| Annual visual examination | At least once a year, US supply | statutory 21 CFR 211.170(b) [4] |
| Retention sample numbering series | Site-defined, unique, traceable | site policy traceability duty at Annex 19 2.4 [2] |
| Batch record retention (for cross-reference) | At least 1 year after batch expiry | statutory 21 CFR 211.180(a) [5] |
Rows badged site policy are internal convention, not a compendial or statutory requirement. Verify every statutory row against the edition in force at your site before use.
7. Precautions
- Never destroy a sample from a batch under complaint, recall or investigation, whatever the register says.
- Do not open a retention sample of a fully packaged unit for routine inspection — opening it destroys its value as a specimen of the pack as distributed.
- Where the site supplies more than one market, apply the longest applicable period and the largest applicable quantity, and state in the SOP which instrument drives each.
- Segregate retention samples of products with different labelled storage conditions rather than averaging a single set point.
8. Deviation handling
Record any shortfall in retained quantity, any storage excursion, any sample destroyed early and any sample that cannot be located as a deviation. Assess the impact on the site’s ability to investigate a complaint for the remainder of the shelf life, notify Head QA, and where a batch is on the market consider whether the deviation affects its continued supply.
9. Annexure-I: Retention sample register
| S. No. | Date drawn | Product / material | Batch No. | Mfg date | Exp date | Batch size | Qty retained | Sample No. | Drawn by | Disposal due | Destroyed on | Verified by |
|---|---|---|---|---|---|---|---|---|---|---|---|---|
10. Annexure-II: Destruction record
Product / material, batch number, retention sample number, quantity destroyed, method of destruction, date, authorised by (Head QA), performed by, witnessed by.
11. Revision history
| Version | Effective | Change | Reason |
|---|---|---|---|
| 00 | DD-MMM-YYYY | First issue | New procedure |
| 01 | DD-MMM-YYYY | Retention periods aligned to revised Schedule M clause 19.8.2 | G.S.R. 922(E) dated 28-Dec-2023 |
Reconciling one SOP against three instruments
Sites supplying more than one market usually discover the mismatch during an audit rather than before it — a quantity sized to a US carve-out that Annex 19 does not allow, or a disposal date calculated from the wrong anchor. A documentation review maps each clause in your procedure to the instrument that actually governs it. See pharmaceutical regulatory compliance consultation for how that review is scoped.
Control sample guidelines: frequently asked questions
Schedule M Part I clause 19.8.2 requires retention samples from each batch of finished product to be kept for at least one year after the expiry date. Samples of active starting materials are kept at least one year beyond the expiry date of the corresponding finished product, and other starting materials other than solvents, gases and water for a minimum of two years if their stability allows.
EU GMP Annex 19 clause 2.1 defines a reference sample as a sample of a batch of starting material, packaging material or finished product stored so it can be analysed if the need arises during the shelf life. A retention sample is a sample of a fully packaged unit of finished product, stored for identification purposes such as presentation, labelling, leaflet, batch number and expiry date. For finished products the two are often identical fully packaged units and may then be regarded as interchangeable.
No. Schedule M clause 19.8.2 requires retention samples to be of a size sufficient to permit at least two full re-examinations, and names no excluded test. EU GMP Annex 19 clause 4.1 likewise excludes no test. The exclusion for sterility and pyrogens comes from 21 CFR 211.170, a United States regulation. Indian SOPs that carry the exclusion without supplying the US market are citing the wrong instrument.
No instrument states one. Schedule M clause 19.8.2 requires storage under the recommended conditions, and Annex 19 clause 5.2 requires storage in accordance with the marketing authorisation. The condition is therefore inherited from each product’s own labelled storage condition. A stated room specification is a site decision derived from the products held in it, and should be documented as site policy rather than as a compendial limit.
Only in one narrow case, and only from 24 September 2026. The revised EU GMP Annex 19 clause 9.5 permits a photographic or digital sample of a re-packaged parallel imported or parallel distributed product where it is duly justified that a physical sample cannot reasonably be retained and this has been agreed in advance with the competent authority. The digital sample must permit full visual examination, be traceable to the batch packaging record, show the safety features and braille information, and comply with Annex 11 for record integrity.
Schedule M Part XII clause 11.6.2 requires reserve samples of each batch of API to be retained for one year after the expiry date assigned by the manufacturer, or three years after distribution of the batch, whichever is longer. For APIs carrying a retest date rather than an expiry date, reserve samples are retained for three years after the batch has been completely distributed. Clause 11.6.3 requires sufficient quantity for at least two full compendial analyses, or two full specification analyses where there is no pharmacopoeial monograph.
Related on Laafon
- Quality control in the pharmaceutical industry — where the retention sample duty sits in the wider QC function.
- Quality assurance in the pharmaceutical industry — the approval and review layer above this procedure.
- Working standard qualification SOP — the other Schedule M sample class most sites document badly.
- Schedule M area requirements by dosage form — sizing the retention area alongside the rest of the plant.
- All SOPs on Laafon — the full QC and QA procedure library.
References
- Ministry of Health and Family Welfare, Government of India. Schedule M — Good Manufacturing Practices and Requirements of Premises, Plant and Equipment for Pharmaceutical Products. Notified by G.S.R. 922(E), 28 December 2023. The Gazette of India: Extraordinary, Part II, Section 3(i). Clauses cited: Part I 19.3(g), 19.6.2, 19.7.2, 19.8.2; Part IV 13.2; Part XII 11.6. Available from: https://cdsco.gov.in/opencms/opencms/en/Notifications/Gazette-Notifications/. Accessed September 2026.
- European Commission. EudraLex Volume 4, EU Guidelines to Good Manufacturing Practice, Annex 19: Reference and Retention Samples. Brussels; 14 December 2005, in operation 1 June 2006. Available from: https://health.ec.europa.eu/system/files/2016-11/2005_12_14_annex19_en_0.pdf. Accessed September 2026.
- European Commission. EudraLex Volume 4, Annex 19: Reference and Retention Samples (revised). C(2026) 4135 final. Brussels; 23 June 2026. Available from: https://health.ec.europa.eu/document/download/831d392e-bcc4-4552-8011-2b724b8655de_en. Accessed September 2026.
- US Food and Drug Administration. 21 CFR 211.170 — Reserve samples. Code of Federal Regulations, Title 21. As amended at 89 FR 51770, 18 June 2024. Available from: https://www.ecfr.gov/current/title-21/chapter-I/subchapter-C/part-211/subpart-I/section-211.170. Accessed September 2026.
- US Food and Drug Administration. 21 CFR 211.180 — Records and reports: general requirements. Code of Federal Regulations, Title 21. Available from: https://www.ecfr.gov/current/title-21/chapter-I/subchapter-C/part-211/subpart-J/section-211.180. Accessed September 2026.
- Saranadasa H. The square root of N plus one sampling rule: how much confidence do we have? Pharmaceutical Technology. 2003 May;27(5):50-62. Reproduced in FDA docket FDA-2017-P-6967-0003. Available from: https://downloads.regulations.gov/FDA-2017-P-6967-0003/attachment_1.pdf. Accessed September 2026.
- European Commission. EudraLex Volume 4: Reference and Retention Samples (revised, applicable as of 24 September 2026). Public Health latest updates; 24 June 2026. Available from: https://health.ec.europa.eu/latest-updates/eudralex-volume-4-reference-and-retention-samples-revised-applicable-24-september-2026-2026-06-24_en. Accessed September 2026.
This procedure is a template for adaptation. It requires local qualification, validation and Quality Assurance approval before use, and every acceptance criterion must be verified against the edition of the instrument in force at your site. Pharmacopoeial texts, EU GMP annexes and Indian statutory instruments change between editions — EU GMP Annex 19 itself changes on 24 September 2026. Technical and educational content only; not medical, legal or investment advice.